 HIPAA Compliant
 ISO 27001 Certified

How to Register a Clinical Trial in the U.S. (2026 Guide for Sponsors)
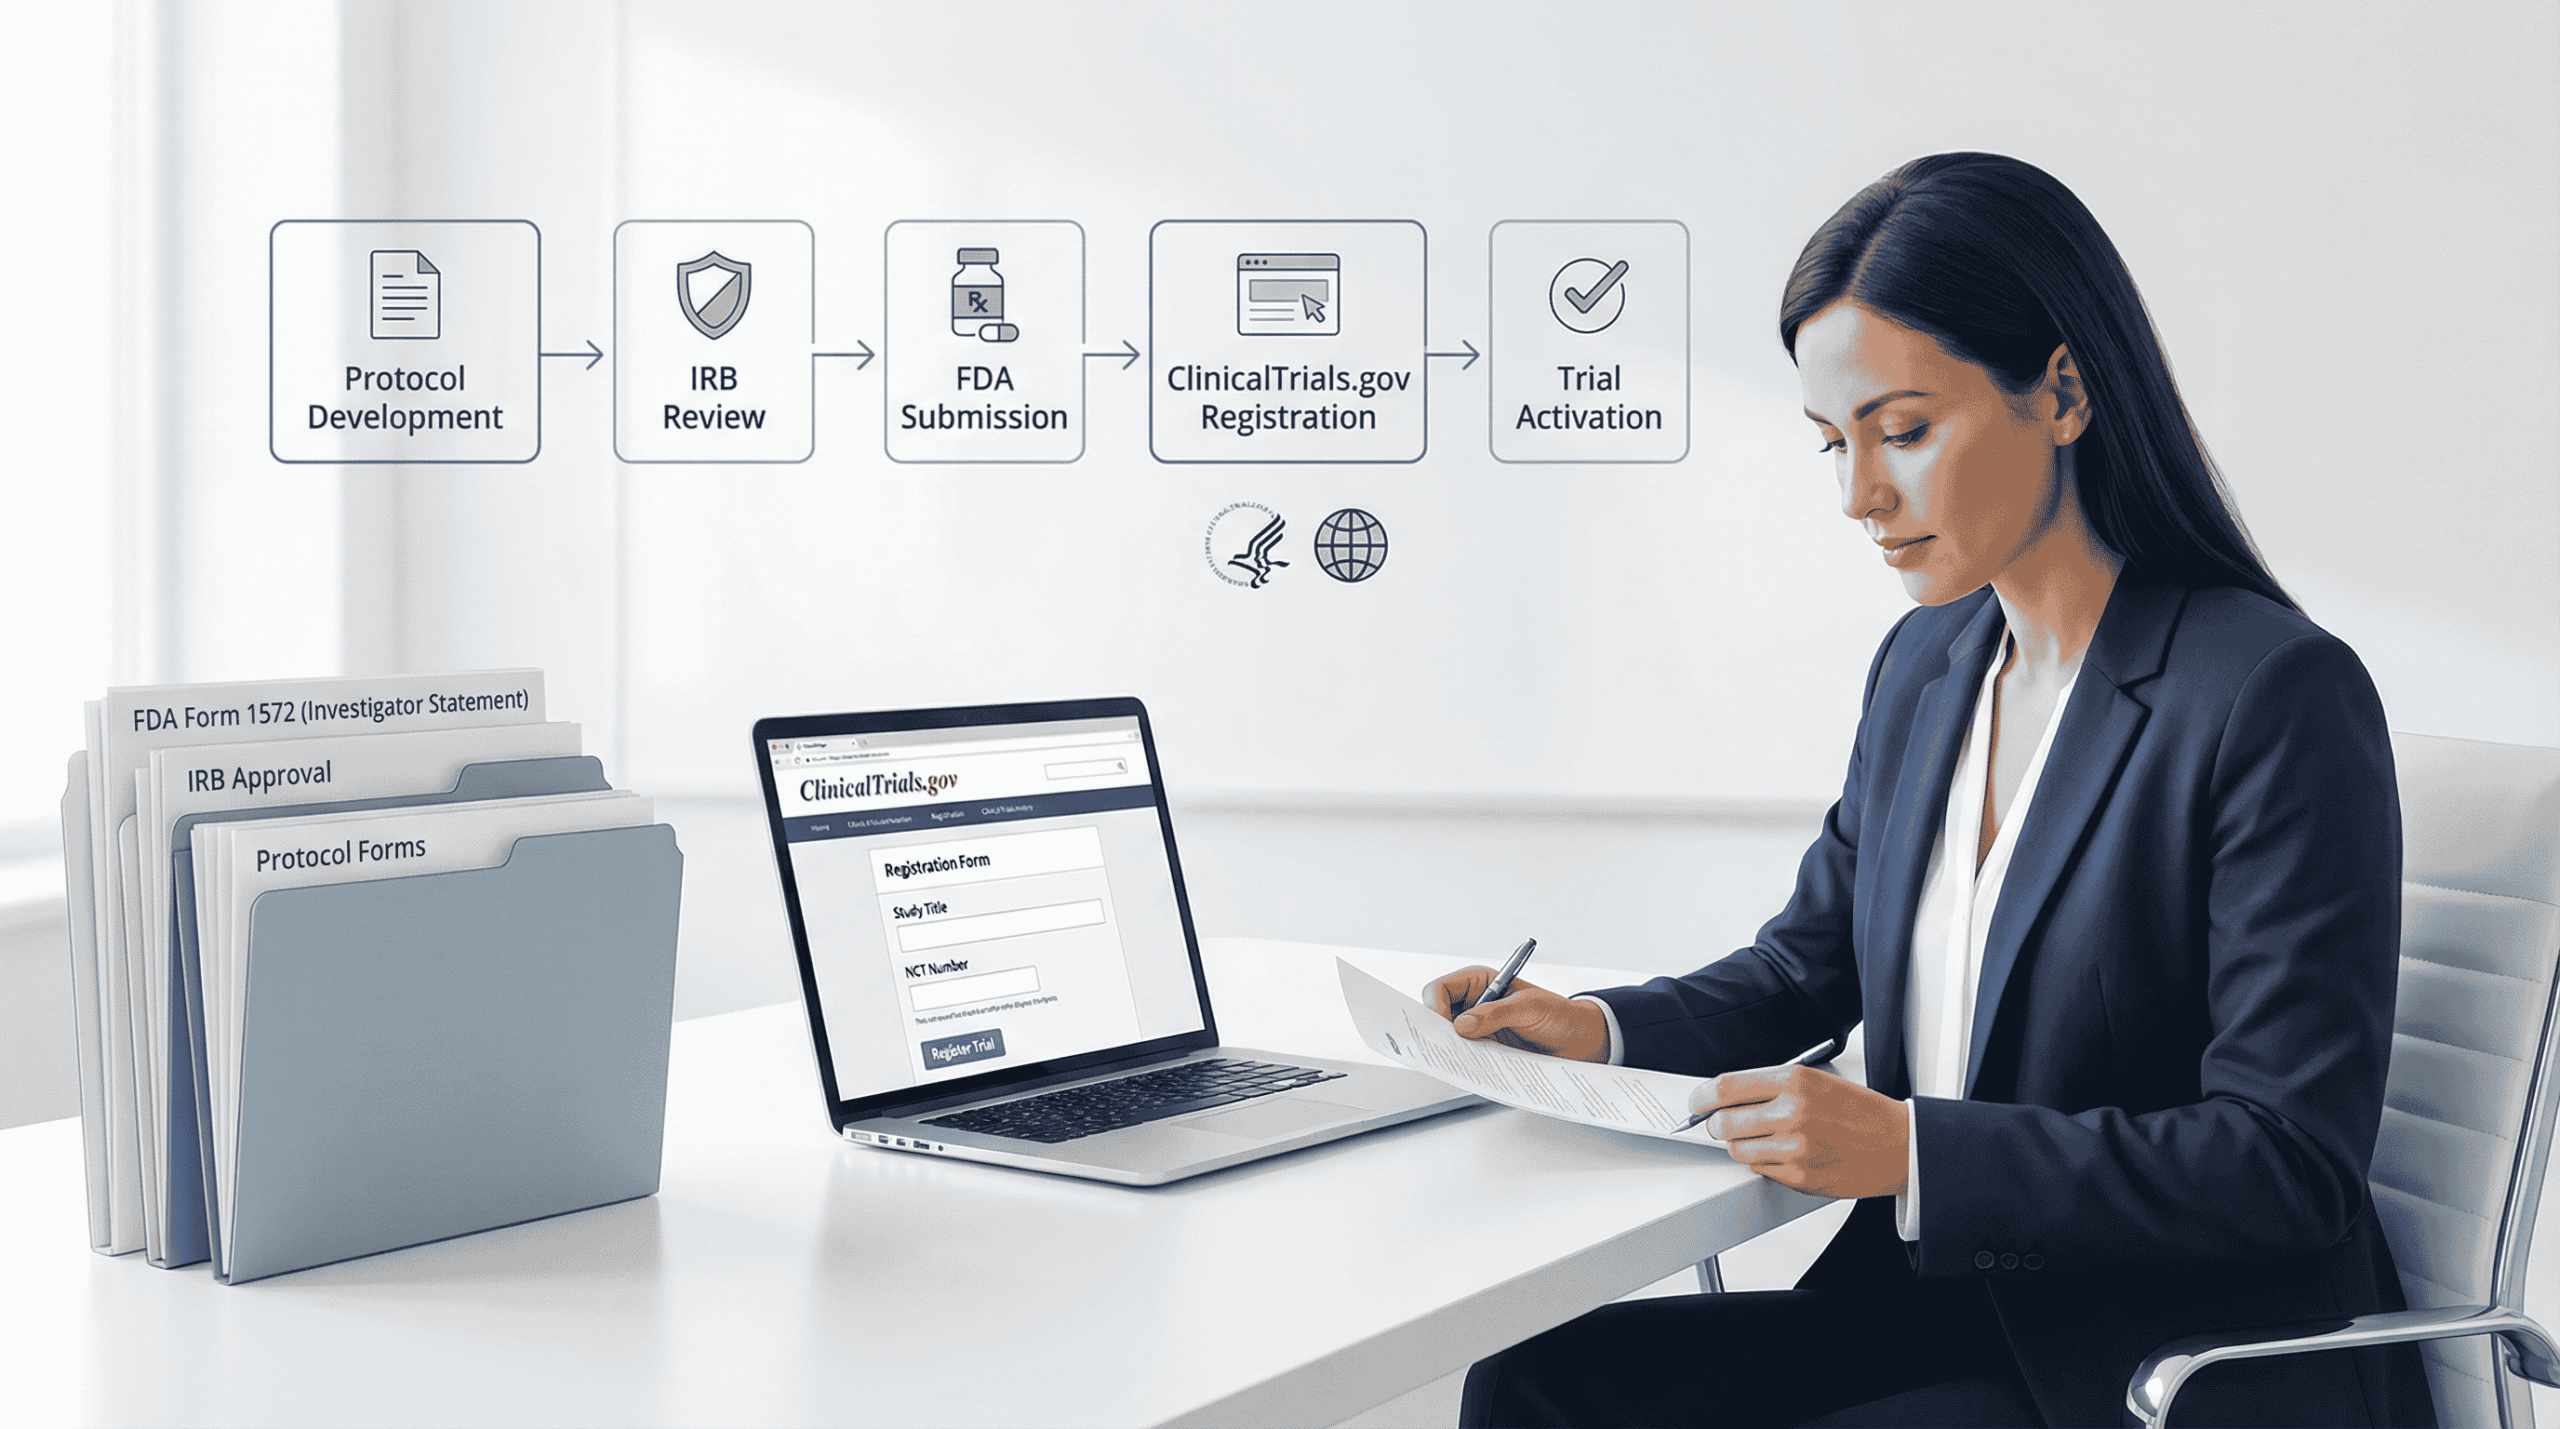
Registering a clinical trial in the United States can feel complicated, especially if your team is juggling IND or IDE decisions, IRB timelines, and ClinicalTrials.gov requirements at the same time. The good news is that the process becomes far more manageable once you understand the sequence and what information each step requires.
This guide explains the registration process in simple terms so sponsors and research teams can avoid delays and keep the study startup on track.
(Note: This guide is informational and does not replace legal or regulatory advice. Requirements evolve, and Sponsors should always verify current regulations.)
Key Takeaways
You will learn:
How DecenTrialz supports pre screening and early readiness
When your study needs an IND or IDE
What documents IRBs expect
What information ClinicalTrials.gov requires in 2026
How PRS review works
Where sponsors often lose time
Step 1: Determine Whether Your Study Requires FDA IND Registration
Before planning recruitment or registration, Sponsors must determine whether an investigational product requires an Investigational New Drug (IND) application.
What an IND Is?
An IND is FDA’s mechanism for overseeing the safety of clinical investigations involving drugs and biologics that are new, used in new ways, or used in new combinations. The IND gives FDA the opportunity to review your plan before people are exposed to the product.
When an IND Is Required
Generally, an IND is needed when your study:
- Involves a new, unapproved drug or biologic
- Uses an approved product for a new indication
- Changes route, dose, or regimen in a way that may increase risk
- Tests new combinations of approved products
Some studies (for example, certain observational or non-interventional drug studies) may be exempt, but Sponsors should document the rationale.
Types of INDs
- Commercial IND – submitted by a company developing a product for marketing
- Investigator IND – submitted by an individual investigator (often academic)
- Emergency Use IND – for urgent situations where treatment cannot wait
Documents Typically Needed
- Protocol
- Investigator brochure
- Preclinical safety data
- Chemistry, manufacturing, and controls (CMC) details
- Informed consent templates
- Safety monitoring and reporting plan
The 30-Day IND Review Period
Once FDA receives the IND, there is a 30-day statutory review period. During this time, the Sponsor must wait 30 calendar days before initiating the study. If FDA does not place the IND on clinical hold, the IND automatically goes into effect at the end of the 30 days, or earlier if FDA explicitly gives the go-ahead.
Enrollment may begin only after:
- The IND is in effect (no clinical hold), and
- IRB approval has been obtained.
Device Studies: Brief IDE Decision Framework
If your study involves a device rather than a drug/biologic:
- Significant risk (SR) devices generally require an FDA Investigational Device Exemption (IDE) approvalplus IRB approval before starting.
- Non-significant risk (NSR) devices do not require a full IDE submission to FDA; they follow “abbreviated IDE” requirements and only need IRB approval, but must still meet FDA device regulations.
- Some device studies are exempt when using approved devices within labeling or under specific regulatory criteria.
Clarifying drug vs. device pathways early helps align the registration clinical trial sequence and documentation.
Common Sponsor Mistakes
- Assuming IND or IDE is “approved” rather than understanding the automatic-effect or “deemed approved” frameworks
- Submitting incomplete safety or CMC information
- Vague primary outcomes and unclear risk justification
Failing to document why an IND/IDE is or is not required
Step 2: Prepare for IRB Review — IRB Approval Steps
An Institutional Review Board (IRB) protects participant rights and welfare. IRB approval is required in addition to FDA oversight before enrollment may begin.
What an IRB Is
An IRB is an independent ethics committee that reviews clinical research to ensure risks are minimized and reasonable, consent is understandable, and privacy protections are in place.
Core IRB Submission Package
Typical documents include:
- Full protocol
- Informed consent and (if applicable) assent forms
- Recruitment materials and scripts
- Investigator brochure or device manual
- Safety information and risk-benefit description
- Data protection and confidentiality plans
Central vs. Local IRB
- Central IRB: Often used for multi-site industry-sponsored studies, providing a single, consistent review.
- Local IRB: Common at academic centers; may require institutional templates or policies.
Timing and Sequencing With IND/IDE
IRBs may review protocols while FDA is reviewing an IND or IDE, but enrollment cannot start until:
- The IND is in effect (or IDE is approved or deemed approved for NSR devices), and
- IRB approval is in place.
Many IRBs expect documentation of the IND or IDE status before final approval is implemented, even if review begins in parallel.
Common Reasons for IRB Delays
- Consent forms lacking key risk or contact information
- Inconsistent details between protocol, consent, and recruitment materials
- Unclear data-handling or monitoring plans
- Missing device risk rationale (SR vs. NSR)
- Repeated minor wording issues that slow approval cycles
Additional Requirement: Informed Consent Posting (Common Rule)
For certain HHS-supported clinical trials, one IRB-approved consent form used to enroll participants must be posted on a designated federal website (such as ClinicalTrials.gov) after recruitment closes and no later than 60 days after the last study visit.
Sponsors should plan this posting as part of their documentation timeline.
Step 3: Register the Trial on ClinicalTrials.gov
ClinicalTrials.gov is the primary public registry for U.S. clinical trials and the focus of many ClinicalTrials.gov guide resources.
Who Must Register?
There are three overlapping policy drivers:
- FDAAA 801/ Final Rule (HHS): Requires registration of “Applicable Clinical Trials” (ACTs), generally Phase 2–4 drug and biologic trials and certain device trials; Phase 1 drug trials are typically exempt.
- NIH Policy: Requires registration and results reporting for all NIH-funded clinical trials, regardless of phase, intervention type, or whether they are ACTs.
- ICMJE Policy: Requires prospective registration of all “clinically directive” trials (any trial assigning human subjects to an intervention to study health outcomes) as a condition for publication.
This means a Phase 1 drug trial might be exempt from FDAAA registration but still must register if NIH-funded or if publication in an ICMJE-aligned journal is planned.
When Registration Is Required
- FDAAA 801 / NIH policy: No later than 21 days after enrollment of the first participant for applicable and NIH-funded trials.
- ICMJE: Requires prospective registration—before the first participant is enrolled.
Sponsors intending to publish or follow common institutional standards should treat “before first enrollment” as the practical default for clinical trial registration USA.
Key Information Needed for Registration
Before using PRS, gather:
- Official study title
- Brief summary (plain language, non-promotional)
- Study design (phase for drugs, or feasibility/IDE terminology for devices; allocation, masking, primary purpose)
- Eligibility criteria (clear inclusion and exclusion)
- Outcome measures with specific time frames
- Study arms and interventions
- Enrollment type and target sample size
- Facility locations
- Oversight details (IND/IDE status, FDA regulated product flags)
- Sponsor and responsible party contacts
Phase labels (Phase 1, 2, 3, 4) apply primarily to drug and biologic studies; device trials may use alternative descriptors (for example, feasibility, pivotal).
Step 4:Using the Protocol Registration and Results System (PRS)
The PRS is the web-based system that manages protocol registration and results submission.
How PRS Works
- Set up an organizational account – A PRS administrator at the institution or Sponsor manages user access.
- Enter structured data elements– Using standardized fields for design, outcomes, eligibility, locations, and oversight.
- Release the record for Quality Control (QC) review – ClinicalTrials.gov staff review for clarity and completeness.
Major vs. Advisory QC Issues
Updated PRS procedures distinguish:
- Major Issues: Must be corrected or addressed to meet QC criteria.
- Advisory Issues: Suggestions to improve clarity; not strictly required but recommended.
For results, ClinicalTrials.gov may post records within 30 days of submission even if QC review is not complete, with notations that QC review is ongoing.
Common PRS Problems to Avoid
- Outcome measures without specified time frames
- Vague or overlapping primary and secondary endpoints
- Missing locations or facility information
- Inconsistent recruitment status vs. actual site activity
- Overuse of abbreviations without explanation
A concise internal checklist can prevent repeated QC cycles and keep your registration clinical trial timeline on track.
Step 5: Understand Registration Timelines
Thinking about sequence early prevents regulatory surprises.
Typical Operational Sequence
- Determine regulatory pathway (IND or IDE; drug vs. device).
- Submit IND or IDE to FDA.
- Wait 30 days for IND to go into effect (or for IDE approval/NSR determination), unless FDA allows an earlier start.
- Obtain IRB approval.
- Register the trial on ClinicalTrials.gov using PRS.
- Begin enrollment only after IND/IDE is active and IRB approval is in place.
Registration Deadlines You Must Balance
- FDAAA / NIH: Register within 21 days after the first participant is enrolled for ACTs and NIH-funded trials.
- ICMJE: Register before the first participant is enrolled to remain eligible for publication.
For most Sponsors, the safest operational approach is to complete ClinicalTrials.gov registration before first enrollment, even if the law allows a 21-day window.
Consequences of Delayed or Inaccurate Registration
- Civil monetary penalties under FDAAA 801
- Potential withholding of NIH funds
- Ineligibility for publication in ICMJE-aligned journals
- Reputational risk and reduced public trust
Step 6: Common Errors Sponsors Should Avoid
Across drug, biologic, and device trials, Sponsors frequently encounter similar problems:
- Missing or non-specific primary outcome measures
- Outcome measures without time frames
- Ambiguous eligibility criteria that do not match the protocol
- Incorrect or inconsistent phase or study description, especially for device trials
- Failure to list all active locations or update when sites open or close
- Out-of-date recruitment status (e.g., shows “Recruiting” when enrollment is closed)
- Protocol amendments not reflected in the ClinicalTrials.gov record
- Not posting required consent forms for certain federally supported trials
Reviewing these items as part of a pre-release QC checklist can greatly improve the quality of your ClinicalTrials.gov record.
How DecenTrialz Helps Simplify Trial Readiness
DecenTrialz does not file INDs, IDEs, IRB submissions, or ClinicalTrials.gov registrations. Those responsibilities remain with sponsors.However, a large portion of registration challenges come from inconsistent eligibility criteria, unclear recruitment materials, or misaligned workflows.
DecenTrialz strengthens early trial readiness by helping sponsors:
- Translate protocol eligibility into structured digital pre screeners
- Ensure participant facing materials are consistent
- Reduce mismatches between protocol text and operational workflows
- Organize participant information for cleaner site review
- Deliver only pre qualified participants once recruitment begins
This improves study startup efficiency and helps prevent avoidable screen failures.
Registering a clinical trial in the United States becomes far less stressful when you follow the correct sequence. By confirming IND or IDE requirements early, preparing complete IRB materials, entering accurate information in ClinicalTrials.gov, and avoiding common mistakes, sponsors can protect timelines and demonstrate transparency.
If you want help organizing eligibility criteria, pre screening logic, and early participant workflows before your study launches, DecenTrialz provides a structured and HIPAA compliant way to support your trial readiness.
Learn more:www.decentrialz.com Contact our team:www.decentrialz.com/contact
Was this article helpful?
Similar Blogs
Clinical Trial Patient Matching: Why Do Sponsors Need More Than a Public Registry Listing?
Every applicable clinical trial conducted in the United States must be registered in the f...
 Swaroop ESD
DecenTrialz vs traditional recruitment platforms: how does full-service matching differ from self-service trial search?
Comparisons between DecenTrialz and traditional recruitment platforms usually come down to...
Vamshi Kantoju
RN-led pre-screening in clinical trials: what does nurse review add that AI matching cannot?
Artificial intelligence has changed how quickly clinical trial candidates can be identifie...
Deeksha GittaStay Informed. Stay Connected.
Get updates on verified clinical trials, emerging treatments, and research breakthroughs directly in your inbox. No spam, just science that matters.
